The UK Responsible Person for medical devices with the broadest service list may not be the best fit for your business. What matters is whether the provider’s responsibilities, documentation process and communication arrangements are clearly defined for your devices and target market.
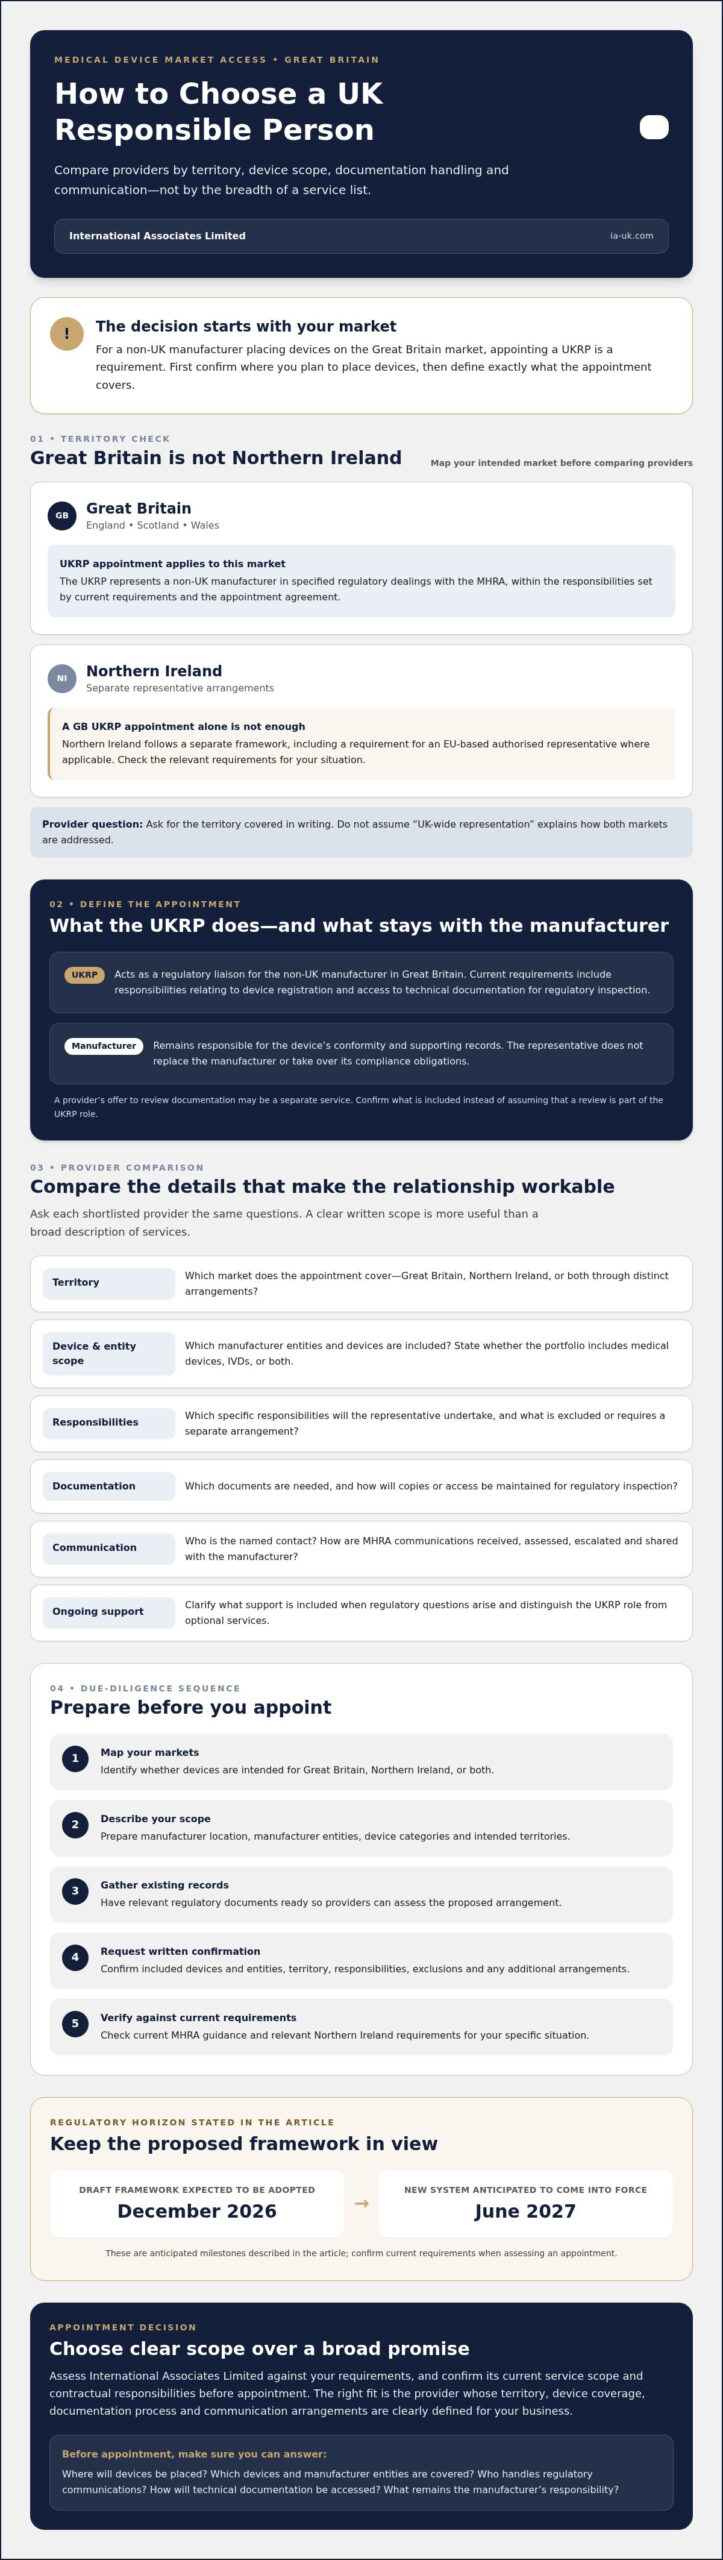
For non-UK manufacturers placing devices on the Great Britain market, appointing a UKRP is a requirement. Yet provider descriptions can make it difficult to distinguish the representative’s role from related services, or to understand what support is included when regulatory questions arise. That uncertainty matters as the UK’s draft medical device framework is expected to be adopted in December 2026, with the new system anticipated to come into force in June 2027.
This guide explains what to confirm before appointing a UK Responsible Person, then compares providers using practical criteria for regulatory scope, communication, technical documentation access and ongoing support. It also explains the distinction between Great Britain and Northern Ireland, so you can assess whether a provider’s stated remit matches the markets where you intend to place your devices. The goal is a clear, workable relationship, not simply a broad service description.
Key Takeaways
- Start by identifying the markets where your device will be placed. Great Britain and Northern Ireland have different representative requirements.
- Compare each UK Responsible Person for medical devices by its confirmed territory and device scope, not by broad service descriptions.
- Ask how technical documentation will be handled, who your named contact is, and how regulatory requests will be escalated.
- Prepare device and manufacturer details, intended territories, and existing regulatory documents before shortlisting providers.
- Assess International Associates Limited against your requirements, and confirm its current service scope and contractual responsibilities before appointment.
Choosing a UK Responsible Person for Medical Devices: Start with Your Market Scope
Identify where you intend to place the device before comparing providers. The UK Responsible Person for medical devices is a regulatory representative appointed by a manufacturer based outside the UK for the Great Britain market, comprising England, Scotland and Wales. In this role, the UKRP acts as a regulatory contact with the Medicines and Healthcare products Regulatory Agency (MHRA), within the responsibilities set by current requirements and the appointment agreement.
In Great Britain, the UKRP represents a non-UK manufacturer in specified regulatory dealings. It does not replace the manufacturer or transfer the manufacturer’s obligations. The manufacturer remains responsible for its device’s conformity and supporting records. An EU Authorised Representative is a distinct role under EU requirements, not another name for a UKRP. A company may offer both services, but confirm the territory and contractual scope of each appointment separately.
Which territory does a UK Responsible Person cover?
Map your launch plans to Great Britain, Northern Ireland, or both. These markets do not have identical representative arrangements: a UKRP appointment applies to Great Britain and is not, by itself, a route to meeting Northern Ireland’s requirements. Northern Ireland follows a separate framework, including a requirement for an EU-based authorised representative where applicable. Check current MHRA guidance and relevant Northern Ireland requirements before appointing a provider, particularly if your distribution plans span both territories.
Ask each provider to state in writing which territory its proposed service covers. Don’t rely on phrases such as “UK-wide representation” unless the provider explains how the arrangement addresses each market.
Which devices and manufacturers need representation?
Before requesting proposals, record the manufacturer’s location and identify the device categories in scope, including whether your portfolio includes medical devices, in vitro diagnostic devices (IVDs), or both. Under current Great Britain arrangements, non-UK manufacturers must appoint a UKRP before placing devices on that market. Confirm how the requirement applies to your specific situation rather than assuming every device or manufacturer has an identical arrangement.
Give shortlisted providers enough information to assess the proposed scope, then request written confirmation of:
- the devices and manufacturer entities included;
- the territory covered by the appointment;
- the representative’s specified responsibilities; and
- any exclusions or additional arrangements you may need.
This initial scope check makes provider comparisons more meaningful and helps prevent a Great Britain service description from being mistaken for coverage across every UK territory.
What a UK Responsible Person Does and What the Manufacturer Still Owns
A UKRP appointment creates a defined representative relationship. It doesn’t make the representative the manufacturer or transfer the manufacturer’s compliance responsibilities. The official government guidance describes the Great Britain requirements, including the appointment of a UK Responsible Person by manufacturers based outside the UK. Use current MHRA guidance, rather than a provider’s general service description, to check which duties apply to your device and arrangement.
For Great Britain, the UKRP acts as a regulatory liaison for the non-UK manufacturer. Current requirements include responsibilities relating to device registration and access to technical documentation for regulatory inspection. Verify the precise scope against the applicable rules and reflect it in the written appointment. A provider’s offer to review documentation may be a separate service. Don’t treat it as proof that the UKRP role itself includes a particular review or assessment.
Responsibilities to confirm in the proposed arrangement
Ask the provider to specify which MHRA communications it will receive, who will assess and escalate them, and how the manufacturer will be kept informed. Confirm which documents the provider needs, how copies or access will be maintained, and how updates will be shared when the manufacturer revises records. These arrangements should support timely, controlled responses while making clear who is responsible for each action.
Also ask the provider to identify any registration, reporting, or record-related duties it considers part of its role. Check those statements against current MHRA guidance and the appointment terms. A responsibility matrix can distinguish the provider’s tasks from information and decisions that must come from the manufacturer.
Responsibilities that remain with the manufacturer
Appointing a UKRP provides representation; it does not transfer the manufacturer’s responsibility for device conformity or the evidence supporting it. The manufacturer should retain control of its technical documentation, quality records, and decisions about device changes, while ensuring the representative receives the information needed for its agreed role. For example, if a device or its supporting records change, establish who assesses the change, who updates controlled documents, and how the UKRP receives the relevant current information.
Keep the role distinct from related activities. Technical documentation review and ISO 13485 certification are not substitutes for UKRP representation, and neither changes the manufacturer’s accountability. Manufacturers assessing their quality-system framework can consult an ISO management system certification resource, while confirming separately what their UKRP appointment covers.
International Associates Limited lists EU and UK Authorised Representative services. If you’re assessing its UKRP offering, confirm the current device scope, responsibilities, and documentation arrangements directly, and compare them with your verified requirements. Its regulatory compliance services provide a starting point for reviewing the relevant service information.
How to Compare UKRP Providers: Scope, Documentation, and Responsiveness
Compare providers against the same questions, using written answers rather than broad claims of “global regulatory support.” The right UK Responsible Person for medical devices should be able to explain how its proposed service fits your territories, device portfolio, documentation needs, and communication process. Use a consistent checklist to make differences visible.
| Criterion | What to confirm | Useful evidence |
|---|---|---|
| Territory | Which markets are covered, and what is excluded? | A written scope naming each territory and any separate arrangements needed. |
| Device scope | Which devices, categories, and manufacturer circumstances are included? | Confirmation based on the device and manufacturer information you provide. |
| Named contacts | Who handles routine queries and who is the alternative contact? | Named roles, contact channels, and a clear process for routing requests. |
| Documentation | What records are required, and is review included in the representative service? | Document list, access method, confidentiality provisions, and change process. |
| Escalation | How are regulatory inquiries received, tracked, assigned, and escalated? | A described workflow showing how the manufacturer is informed and involved. |
| Contract terms | How do notification, renewal, termination, and handover work? | Written terms covering service boundaries, fees, and transition arrangements. |
Questions to ask before appointing a UKRP
Ask providers to confirm in writing which devices, territories, and manufacturer circumstances their proposal covers. Then test the workflow with a practical scenario: if a regulatory request arrives or your portfolio changes, who receives it, records it, contacts your team, and follows up? Ask how updates to contact details or product scope are handled, and what notification, renewal, termination, and handover provisions appear in the agreement. Discuss fees directly with each provider rather than relying on assumed market averages.
Assess medical-device experience through relevant examples, defined processes, and clear answers about the device categories in your portfolio. General claims of regulatory expertise alone don’t establish that a provider’s experience or service scope matches your needs.
How to assess technical documentation support
Confirm whether documentation review is included in the UKRP agreement, available as a separate service, or outside the proposed scope. Ask how access is controlled, how revised versions are identified, how confidential records are protected, and how documents can be retrieved when needed. The answers should distinguish representative functions from any additional review service, so responsibilities and fees remain clear.
For broader context on the listed regulatory compliance services, assess each offering separately against your specific requirements.
How to Choose and Appoint a UKRP: A Practical Due-Diligence Process
A disciplined appointment process turns provider proposals into comparable commitments. Use the same device and market information for each enquiry, then verify the proposed scope and working arrangements before signing.
1. Define market scope. Record whether your plans cover Great Britain, Northern Ireland, or both. Confirm current requirements for each intended territory rather than assuming one appointment covers every market.
2. Set out documentation needs. Identify what regulatory documents are available, where controlled versions are held, and how the representative will access records required for its agreed role.
3. Shortlist providers. Compare relevant medical-device experience, stated territory and device coverage, named contacts, and the provider’s process for regulatory communications. Ask what further information it needs to assess your situation.
4. Verify service scope. Request written confirmation of included manufacturers, devices, territories, representative tasks, exclusions, and any separate documentation review services.
5. Review the agreement. Check responsibilities, communication routes, fees, document access, confidentiality, change notifications, termination, and transition arrangements before appointment.
What to prepare before requesting proposals
Prepare a concise scope pack so providers can assess the same facts. Include device categories, manufacturer location and entity details, intended markets, current regulatory status, existing regulatory documents, and technical documentation availability. Identify the internal contacts responsible for regulatory questions and records. Note anticipated portfolio changes, such as planned device additions, and ask each provider what else it needs before confirming coverage.
What to record in the appointment agreement
Ensure the agreement names the covered territories and devices and defines the service boundaries. It should also make clear who receives regulatory communications, who contacts your team, how requests are tracked and escalated, and how document access and updates are managed. Confirm confidentiality provisions, fees, renewal and termination terms, and the process for transferring relevant information if the appointment ends.
Before approval, separate what is established from what still needs confirmation. This prevents assumptions from becoming part of the operational plan.
- Confirm against your own records: manufacturer identity and location, device list, intended territories, available documents, and internal contacts.
- Obtain from the provider in writing: included devices and markets, service boundaries, communication workflow, document handling, fees, and contract terms.
- Verify against current regulatory requirements: whether the proposed arrangement is appropriate for each territory and device category.
To discuss a potential appointment, review UK regulatory compliance support and confirm the current service scope and contractual responsibilities directly.
When International Associates May Suit Your UKRP Requirements
International Associates Limited is one provider to assess if its stated services align with your device portfolio, intended markets, and required level of support. The company lists EU and UK Authorised Representative services, alongside technical documentation review for medical devices and ISO 13485 certification. These are distinct services, so confirm exactly which are included in any proposed UKRP arrangement.
Assessing fit against your device and market plans
Before seeking a proposal, prepare the device categories you need covered, the manufacturer’s location, and the territories where you plan to place devices. Explain whether you’re seeking UK representation alone or considering arrangements for more than one market. This gives the provider the information to clarify whether its current service scope matches your requirements, rather than leaving you to rely on a general description of representative services.
Suitability depends on the provider’s confirmed scope and the appointment terms. Ask it to specify which devices and territories are covered, what responsibilities it will undertake, how regulatory communications will be handled, and what documentation arrangements apply. Confirm any exclusions or separate services in writing before making a decision.
Technical documentation review may be relevant to your records, but it should not be assumed to be part of UKRP representation. ISO 13485 certification is also a separate quality-system service, not a substitute for appointing a representative. Keep these scopes distinct when comparing proposals and assigning internal responsibilities.
Next steps for discussing UK representation
Prepare concise manufacturer and device information before contacting a provider. Include device categories, intended territories, available regulatory documents, and the support you’re seeking. Then request written clarification of covered devices, applicable territories, representative responsibilities, documentation access, communication arrangements, fees, and contractual terms. If your plans include Great Britain and Northern Ireland, ask how the proposed arrangements address each territory rather than assuming one appointment covers both.
International Associates has a Glasgow head office and offices across Europe, Asia, and the Middle East, which may be relevant context for organisations operating across markets. However, a network’s presence does not establish that a particular territory or device is included in a service. Verify the scope that applies to your requirements.
If you’re assessing International Associates as a UK Responsible Person for medical devices, discuss your UK regulatory compliance requirements and request confirmation of the current service scope, device coverage, and contractual responsibilities. This gives you a clear basis for deciding whether the proposed arrangement fits your market plans.
Make Your UKRP Appointment with Clear Scope and Confidence
Selecting a UK Responsible Person for medical devices is a decision about defined responsibilities, not simply a provider’s breadth of services. Confirm the intended territories and device coverage, then agree how regulatory communications, documentation access, and ongoing changes will be managed. Keep the manufacturer’s own compliance responsibilities distinct from the representative’s contracted role.
International Associates lists EU and UK Authorised Representative services, as well as technical documentation review for medical devices as a separate service. Before proceeding, confirm the current scope, applicable territories, responsibilities, documentation arrangements, and fees in writing. This gives both parties a clear basis for an effective working relationship.
To discuss your requirements and clarify whether the listed service may fit your plans, Discuss your UK Responsible Person requirements.
Frequently Asked Questions
Does a non-UK medical-device manufacturer need a UK Responsible Person?
Generally, a manufacturer based outside the UK must appoint a UK Responsible Person before placing a medical device or IVD on the Great Britain market. Requirements depend on the device and intended territory, so confirm how the current rules apply to your product. Northern Ireland follows a different regulatory arrangement, and a UKRP appointment alone doesn’t meet its requirements. Check current MHRA guidance before making market-entry decisions.
What does a UK Responsible Person do for a medical-device manufacturer?
A UK Responsible Person represents a non-UK manufacturer in specified regulatory matters for the Great Britain market. The role includes responsibilities relating to device registration and making technical documentation available for regulatory inspection, as required by applicable rules. The manufacturer remains responsible for device conformity and its supporting records. Confirm the UKRP’s specific duties against current requirements and the appointment agreement, rather than relying solely on a provider’s general service description.
Is a UK Responsible Person the same as an EU Authorised Representative?
No. A UKRP and an EU Authorised Representative serve different regulatory arrangements and territories. A UKRP appointment relates to Great Britain; an EU Authorised Representative addresses applicable EU requirements. Northern Ireland has distinct arrangements, including an EU-based authorised representative where required. Don’t assume one appointment covers Great Britain, Northern Ireland, and EU markets. Confirm the relevant requirements and proposed scope separately for each territory where you intend to place devices.
Can one UK Responsible Person cover all medical-device types?
Don’t assume a provider’s service covers every device type or manufacturer circumstance. Coverage depends on the scope it agrees to provide and the requirements applicable to the device and market. Before appointment, give the provider your device categories and manufacturer details, including whether the portfolio includes IVDs. Ask it to identify included devices, territories, exclusions, and responsibilities in writing, then verify regulatory questions against current guidance.
What documents should I provide to a UK Responsible Person?
The information requested depends on your device, intended market, and agreed service scope. Providers may need manufacturer details, device information, and access to relevant regulatory records, including technical documentation where applicable. Agree which documents are needed and whether copies or controlled access will be used. Also establish how confidentiality is protected, how updated versions are identified, and how the UKRP can retrieve current records if a regulatory request arises.
How do I choose a UK Responsible Person for medical devices?
Compare each provider’s territory and device coverage, defined responsibilities, documentation arrangements, communication process, contract terms, and relevant medical-device experience. Ask who handles regulatory requests, how issues are escalated, and what happens if your product portfolio changes. Obtain written confirmation of the proposed service scope and fees. Before appointment, verify the regulatory details that apply to your device and intended markets against current guidance.
Does appointing a UK Responsible Person transfer the manufacturer’s compliance responsibilities?
No. Representation should not be treated as a transfer of all manufacturer obligations. The manufacturer remains responsible for device conformity and the records supporting it, while the UKRP undertakes the specific representative responsibilities that apply under current rules and the appointment agreement. Define who controls documentation, supplies updates, and responds to requests. Check role boundaries against current regulations and ensure the written agreement reflects the actual arrangement.