Assigning regulatory duties to your European sales distributor might seem like an efficient shortcut, but it creates direct commercial conflicts of interest and compromises control over your proprietary documentation. Choosing the right EU authorised representative for medical devices requires an independent model that isolates legal accountability without limiting your commercial growth.
It’s understandable why non-EU manufacturers feel immense pressure under Medical Device Regulation (EU) 2017/745. Article 11 joint liability has heightened scrutiny from representatives, while mandatory 2026 EUDAMED deadlines and rapid incident reporting protocols demand strict operational precision. You shouldn’t have to sacrifice intellectual property confidentiality or commercial agility simply to satisfy technical file audits.
This guide provides the vital criteria, legal obligations, and strategic frameworks needed to select an independent representative who protects your assets while meeting MDR and IVDR mandates. We examine operational vetting benchmarks, liability structures, and multi-market access strategies to help you make a decisive, legally sound choice.
Key Takeaways
- Appointing an independent regulatory partner rather than a commercial distributor protects proprietary technical documentation and prevents restrictive sales channel conflicts.
- Selecting the right EU authorised representative for medical devices demands rigorous verification of dedicated Person Responsible for Regulatory Compliance (PRRC) staffing and proven auditing infrastructure.
- Contractual mandates must be structured precisely to satisfy Article 11 joint liability requirements under MDR 2017/745 while maintaining complete manufacturer sovereignty.
- Establishing proactive verification protocols ensures smooth execution of mandatory EUDAMED device registrations and incident escalation procedures.
- Consolidating representation across both European Union and United Kingdom frameworks delivers significant administrative efficiencies for multi-market distribution strategies.
Mandatory Role & Legal Exposure: Why Medical Device Exporters Need an EC REP
Non-EU medical device manufacturers cannot legally introduce products into the European Union without an appointed representative based within a member state. Under Regulation (EU) 2017/745 (MDR), appointing a European authorised representative isn’t a mere administrative formality; it’s a mandatory prerequisite for market clearance. The representative’s registered name and physical address must appear on all device labelling, outer packaging, and instructions for use alongside the CE mark. Under the former Medical Device Directives (MDD), representatives operated largely as passive postal contacts. Under the MDR framework, that dynamic has transformed into active, institutional compliance enforcement.
Legal Mandates Under EU MDR 2017/745 Article 11
Article 11 defines the statutory boundary of representative mandates. The representative has an explicit legal duty to verify that the manufacturer has drawn up the EU declaration of conformity and completed an appropriate conformity assessment. Key statutory tasks include:
- Maintaining continuous access to technical documentation and declarations of conformity for at least 10 years after the last device has been placed on the market (15 years for implantable devices).
- Providing European national Competent Authorities with all necessary documentation and device samples upon formal request.
- Informing authorities immediately if a manufacturer acts contrary to MDR obligations, including terminating the mandate if non-compliance persists.
Maintaining these verification layers requires dedicated operational infrastructure, which is why manufacturers rely on structured regulatory compliance protocols to maintain uncompromised oversight.
The Shift in Joint and Several Liability
The most consequential shift introduced by the MDR lies within Article 11(5). Under this clause, an EU authorised representative for medical devices shares direct joint and several liability with the non-EU manufacturer for defective products placed on the market. If a device causes patient harm and the non-EU manufacturer fails to meet general MDR obligations, injured parties can pursue legal compensation directly against the European representative.
Because of this shared exposure, qualified representatives no longer accept mandates without conducting meticulous technical documentation reviews before onboarding. Manufacturers must protect their operations by establishing bilateral contracts that establish strict audit boundaries, define incident escalation timelines, and safeguard core trade secrets during regulatory scrutiny.
Evaluating Representation Models: Independent EC REP vs Commercial Distributor
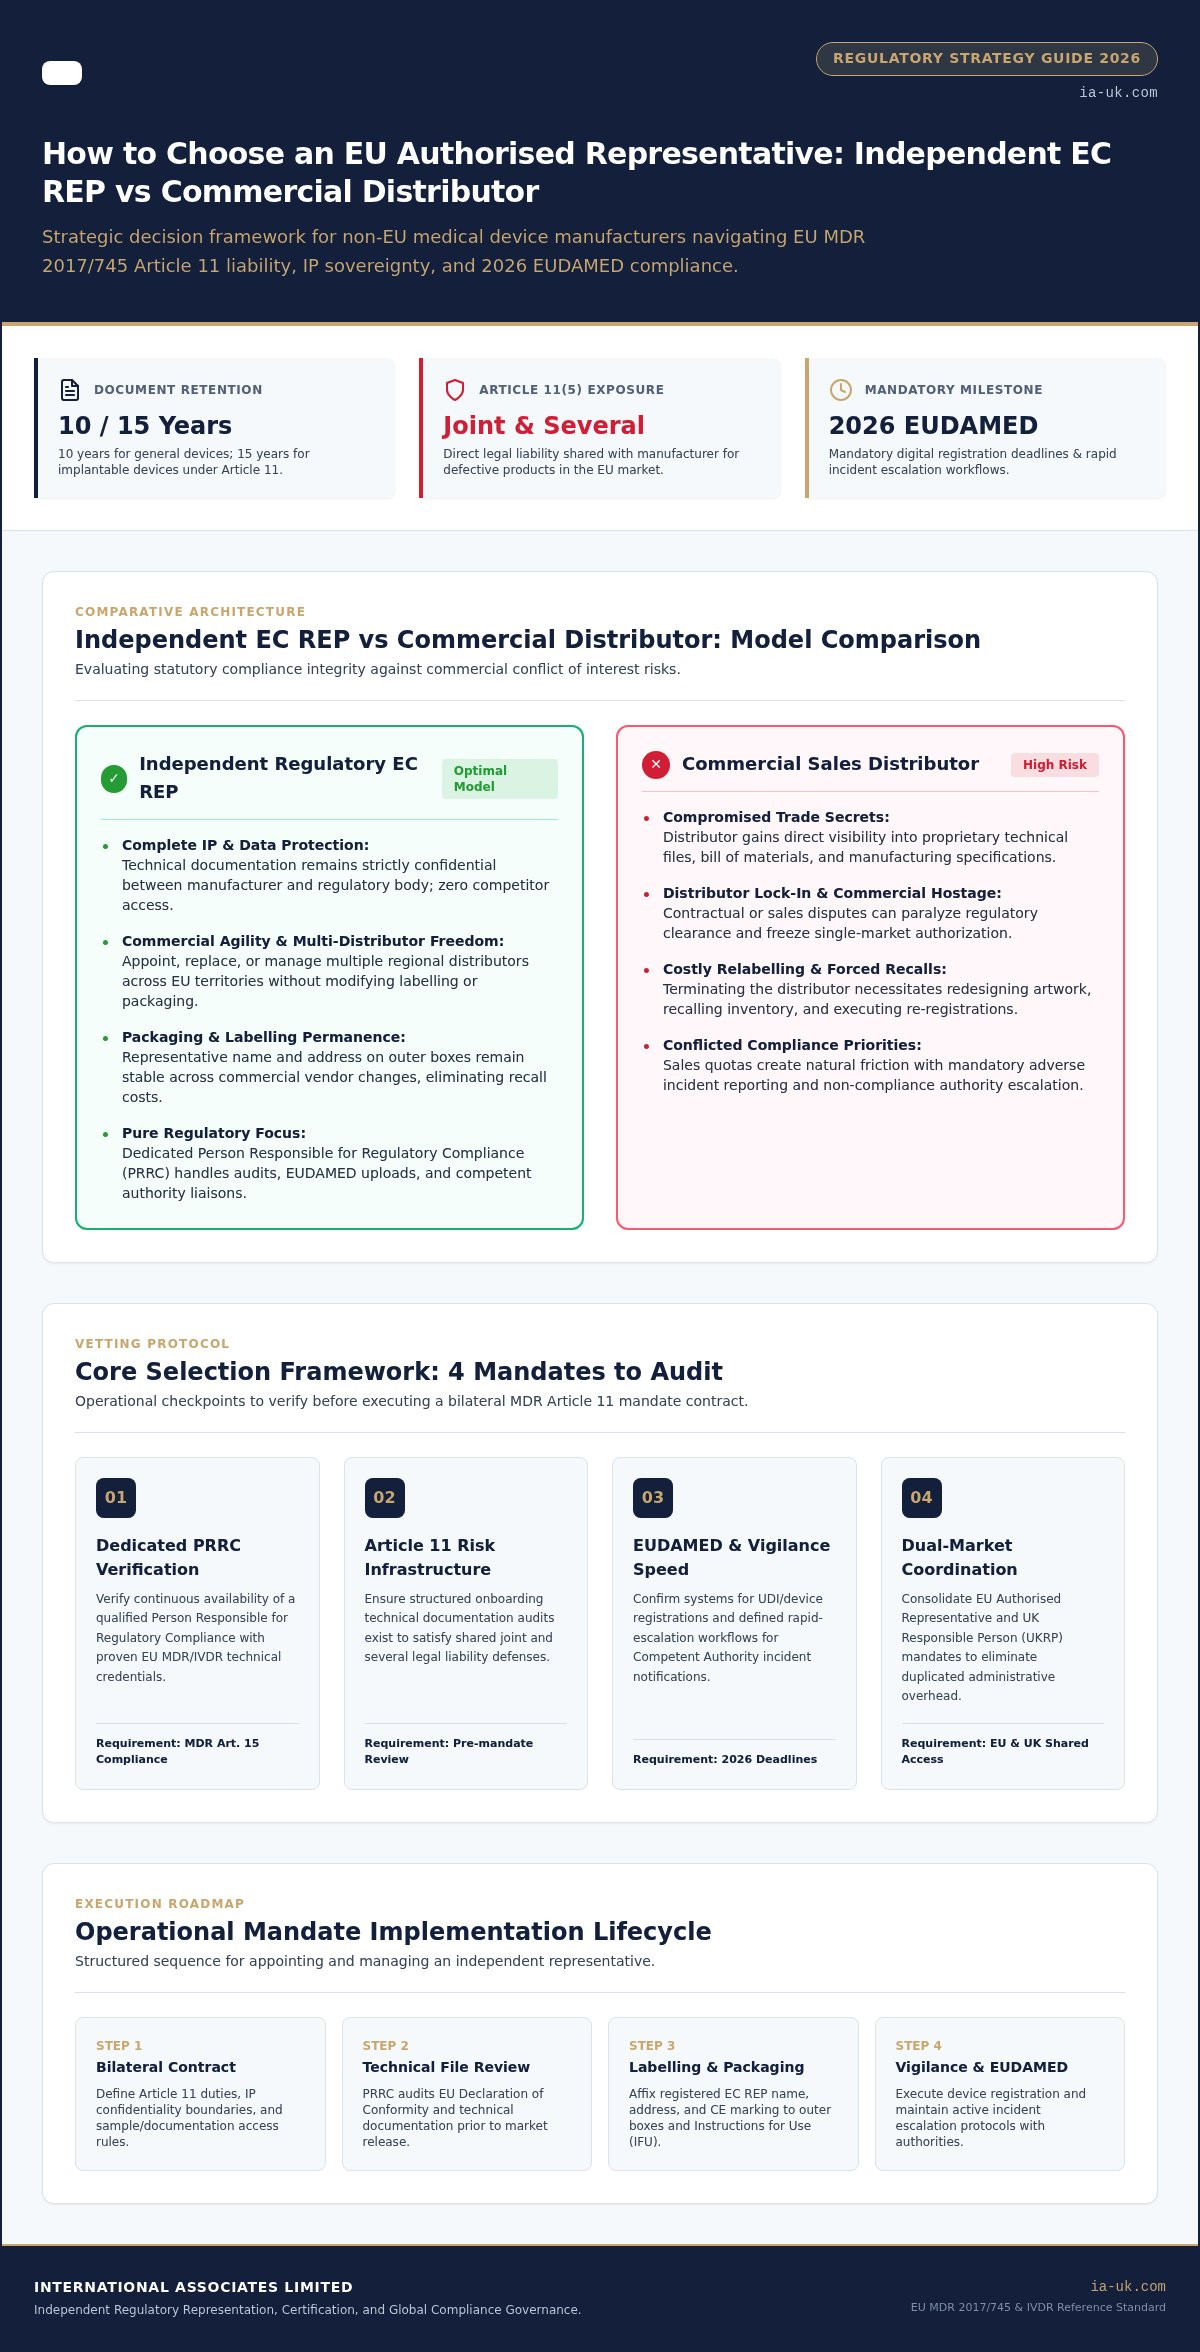
Choosing between an independent compliance body and a commercial distributor is one of the most critical structural decisions a foreign manufacturer faces. While designating a regional sales distributor as your EU authorised representative for medical devices might appear cost-effective initially, it inevitably entangles statutory obligations with commercial sales incentives. Separating legal oversight from territorial sales channels is essential to maintaining long-term market autonomy.
Commercial Conflicts and Supply Chain Vulnerabilities
When an exclusive distributor manages your regulatory mandate, your single-market authorization is tethered directly to a commercial contract. If performance disputes arise or contract negotiations stall, the distributor effectively holds your market access hostage. Because the representative’s identity must appear on all product packaging, terminating that relationship forces costly product recalls, complete label redesigns, and re-registration delays across target territories.
Appointing an independent representative eliminates distributor lock-in. A neutral entity allows you to appoint, manage, or replace multiple regional distributors across different member states without modifying your technical documentation or packaging artwork. This clean operational separation mirrors the strategic structure recommended under the official UK Responsible Person guidance when orchestrating multi-region sales channels across post-Brexit borders.
Protecting Intellectual Property and Technical Files
Statutory compliance under MDR 2017/745 demands that your representative maintains full, direct access to detailed technical files, including clinical evaluations, risk analyses, proprietary manufacturing workflows, and software source data. Granting a commercial distributor unrestricted access to unredacted design dossiers introduces unacceptable intellectual property risks, particularly if that distributor markets competing therapeutic lines or develops proprietary alternatives.
Independent compliance entities function strictly as regulatory fiduciaries under rigid non-disclosure frameworks. By maintaining technical files within an accredited auditing and assurance environment, manufacturers safeguard proprietary clinical intelligence from commercial compromise. Structuring your representation through dedicated regulatory compliance services ensures complete confidentiality while satisfying the demanding verification protocols enforced by European authorities.
Core Selection Criteria for Choosing an EU Authorised Representative
Selecting an EU authorised representative for medical devices requires an exhaustive operational audit rather than a simple price check. Because the representative acts as your regulatory conduit before European authorities, operational incompetence directly threatens your market presence. Manufacturers should evaluate prospective partners against verifiable technical capabilities, internal compliance staffing, and dedicated quality infrastructure.
PRRC Qualifications and Technical Competence
Under Article 15 of Regulation (EU) 2017/745, both the non-EU manufacturer and their representative must have permanent access to a qualified Person Responsible for Regulatory Compliance (PRRC). A single individual cannot legally act as the PRRC for both parties simultaneously. When vetting candidates, confirm that the representative’s PRRC possesses formal credentials matching your device portfolio:
- Documented university degrees in law, medicine, pharmacy, or engineering, accompanied by at least one year of professional regulatory experience in European medical devices.
- Alternatively, a minimum of four years of verifiable professional experience in regulatory affairs or quality management systems covering your device classification.
- Demonstrated technical fluency across complex device categories, including active implantable hardware, sterile consumables, or software as a medical device (SaMD).
EUDAMED Management and Vigilance Response Workflows
Operational execution hinges on secure digital infrastructure. Starting May 28, 2026, the use of EUDAMED’s first four modules became mandatory, requiring all new devices, systems, and procedure packs to be registered before hitting the market. For legacy products already marketed before that date, manufacturers face a strict November 27, 2026 registration deadline.
Your representative must operate automated protocols to map European Medical Device Nomenclature (EMDN) codes and Basic UDI-DI attributes within these centralized systems. Inquire about their formal incident escalation pathways. When serious incidents occur, European market surveillance mandates require reporting within 2 to 15 calendar days depending on severity. Ensure your representative maintains established channels with national Competent Authorities to coordinate Field Safety Corrective Actions (FSCAs) without administrative bottlenecks.
Alignment with ISO 13485 Quality Management Systems
Representative workflows must interface directly with your existing quality architecture. Independent bodies with established capabilities in medical device quality certification deliver unmatched procedural discipline. Selecting a partner experienced in accredited technical reviews and ISO 13485 certification ensures that technical documentation updates, post-market surveillance reports, and vigilance records remain fully harmonized. This structured alignment ensures immediate audit-readiness during sudden unannounced inspections by European Notified Bodies.
Operational Implementation: Appointing and Managing Your Representative
Executing an efficient onboarding process requires a structured operational roadmap. Appointing an EU authorised representative for medical devices follows a clear four-step path: finalising the bilateral mandate, completing preliminary technical documentation verification, generating your Single Registration Number (SRN) via EUDAMED actor registration, and updating device labelling. Clear execution preserves market access and prevents regulatory friction.
Drafting the Article 11 Mandate Agreement
The written mandate forms your legal baseline under MDR Article 11(3). This contract must clearly define verification obligations, sample access terms, and data retention schedules. Crucially, the agreement must establish liability boundaries and manufacturer indemnification clauses without compromising the representative’s statutory oversight. It’s critical to draft explicit terms that safeguard proprietary file access, ensuring your technical dossiers remain protected against broad disclosure while still meeting competent authority inquiries.
Labelling, Packaging, and Administrative Execution
Once the mandate takes effect, physical device packaging must display the standardized “EC REP” symbol adjacent to the representative’s registered legal entity name and address. This requirement applies across outer cartons, sterile barrier packaging, and instructions for use (IFUs). To avoid costly write-offs, manufacturers should establish phased inventory transitions that coordinate print production with supply chain rollouts. Maintaining governance through accredited management system certification providers guarantees that packaging change controls align with relevant traceability standards.
Executing Seamless Representative Transfers (Article 12)
Switching providers requires strict adherence to MDR Article 12. The transition demands a formal tripartite agreement executed between the manufacturer, the outgoing representative, and the incoming entity. This legal document establishes:
- The exact operational date when the outgoing entity’s mandate concludes and the successor assumes statutory responsibility.
- The handover schedule for all technical documentation, complaint histories, and post-market vigilance records.
- Defined inventory run-out allowances that permit existing distributed stock bearing previous labelling to remain on the market without regulatory recall.
A well-executed transfer ensures zero market interruption. If you need a reliable, conflict-free partner to manage your European regulatory presence, you can appoint an independent authorised representative to protect your commercial autonomy today.
Strategic Dual-Market Access: Coordinating EU and UK Compliance
Operating across both the European Union and the post-Brexit British market presents a clear logistical hurdle for foreign medical device manufacturers. While Great Britain operates under the UK Medical Devices Regulations 2002, CE-marked devices compliant with EU MDR can legally remain on the market in England, Scotland, and Wales until June 30, 2030. Even so, placing products across these adjacent territories requires distinct legal entities to satisfy cross-border accountability.
Bridging the Gap: EC REP vs UK Responsible Person (UKRP)
Manufacturers outside the UK must designate a UK Responsible Person (UKRP) to register devices with the MHRA, while non-EU entities require an appointed EU representative for European market clearance. Managing these mandates separately introduces redundant administrative layers. By adopting a harmonized dual-labeling strategy, manufacturers can print both representative markers on their packaging simultaneously:
- The standard “EC REP” symbol with the European entity’s registered address for EU member states.
- The “UKRP” designation alongside the UK entity’s registered address for Great Britain.
- Consolidated vigilance procedures that satisfy both the MHRA incident reporting guidelines and MDR post-market surveillance requirements.
Operating a synchronized strategy eliminates the need for segregated inventories. Manufacturers can gain additional procedural efficiency by consulting technical analyses from established ISO 13485 certification bodies in the UK, ensuring their underlying quality documentation satisfies both authorities concurrently.
Integrated Compliance Delivery with International Associates Limited
International Associates Limited bridges these dual-market demands through unified compliance infrastructure. Operating from headquarters in Glasgow with established representation facilities across the European Union, the organization provides seamless UKRP and EU representation services under a coordinated oversight framework. This dual-market capacity eliminates duplicate file reviews, reduces cross-channel communication friction, and streamlines post-market reporting into a single operational interface.
Selecting International Associates Limited as your EU authorised representative for medical devices guarantees complete commercial independence. Because the organization does not operate sales or logistics networks, your commercial distribution channels remain entirely autonomous. Non-EU manufacturers retain full sovereignty over their intellectual property, distribution agreements, and territorial pricing. To establish an independent compliance framework spanning both European and British jurisdictions, connect with our specialists through the regulatory compliance services portal.
Strengthen Your Market Access with Independent Representation
Navigating MDR 2017/745 and IVDR 2017/746 mandates requires a deliberate separation between commercial sales distribution and statutory regulatory oversight. Appointing an independent compliance partner shields proprietary technical files from market competitors, prevents distributor lock-in, and maintains uncompromised control over regional pricing strategies.
Selecting the right EU authorised representative for medical devices transforms Article 11 obligations from an operational liability into a scalable foundation for global expansion. When legal accountability is isolated within a dedicated compliance entity, your business retains total agility to adjust sales channels across member states without administrative disruption.
You don’t have to compromise commercial flexibility to satisfy European regulatory mandates. With decades of regulatory assurance and auditing leadership operating across global compliance sectors, International Associates provides comprehensive in-house expertise delivering integrated EU AR, UKRP, and ISO 13485 assessment frameworks. Secure Independent EU and UK Representation with International Associates to protect your commercial autonomy and expand with confidence.
Frequently Asked Questions
Can our commercial distributor also act as our EU Authorised Representative?
Yes, a commercial distributor can legally act as your representative, but doing so introduces serious operational and commercial risks. Entangling distribution with regulatory compliance forces you to expose proprietary technical files and creates commercial distributor lock-in. If a sales dispute occurs, switching distributors requires halting shipments and redesigning product labeling. Appointing an independent EU authorised representative for medical devices keeps your sales channels flexible and autonomous.
What are the legal liabilities of an EU Authorised Representative under MDR 2017/745?
Under MDR Article 11(5), the representative shares direct joint and several liability with the non-EU manufacturer for defective devices placed on the EU market. If a non-EU manufacturer fails to compensate an injured patient or fulfill regulatory obligations, European authorities or claimants can pursue the representative directly. Because of this statutory exposure, qualified representatives enforce rigorous technical file audits and post-market surveillance checks before accepting mandates.
Is an EU Authorised Representative required if we already have a UK Responsible Person?
Yes, you still require an EU Authorised Representative because the UK Responsible Person (UKRP) mandate only covers Great Britain under UK regulations. Following Brexit, regulatory jurisdictions are completely separate. An entity based in the UK cannot represent your products within the EU, and an EU-based representative cannot submit registrations directly to the MHRA. Non-EU manufacturers selling in both territories must maintain distinct representation across both jurisdictions.
What credentials must the Representative’s Person Responsible for Regulatory Compliance (PRRC) hold?
Under MDR Article 15, the representative’s PRRC must demonstrate verified expertise in European medical device regulations. This requires a recognized university degree in law, medicine, pharmacy, or engineering, accompanied by at least one year of European regulatory experience. Alternatively, four years of verifiable professional experience in regulatory affairs or quality management systems fulfills the mandate. Your company’s PRRC cannot simultaneously serve as your representative’s PRRC.
How long does it take to complete the transfer to a new EU Authorised Representative?
A standard transfer typically takes between four and twelve weeks, depending on the speed of technical file reviews and contract finalization. The process requires drafting an Article 12 tripartite agreement between the manufacturer, outgoing representative, and incoming entity. Additional time must be allocated to update actor registrations in EUDAMED and align inventory exhaustion schedules so existing distributed stock avoids customs delays or market quarantine.
Does the EU Authorised Representative need to hold a full copy of our technical documentation?
Yes, your representative must maintain continuous, permanent access to a complete copy of your technical documentation and declaration of conformity. Under MDR Article 11(3), they must retain these files and present them to national Competent Authorities upon request for at least 10 years after the last device is placed on the market (15 years for implantables). Secure digital hosting ensures statutory compliance while safeguarding proprietary IP.
Can a non-EU manufacturer appoint multiple EU Authorised Representatives for different devices?
Yes, a manufacturer can appoint different representatives, provided each representative holds the mandate for a distinct generic device group. Under MDR Article 11(2), you cannot appoint more than one representative for the same device category or catalog line. While legally permissible across varied product families, consolidating your portfolio under a single independent EU authorised representative for medical devices minimizes administrative overhead and streamlines EUDAMED registrations.